








GM1 Gangliosidosis
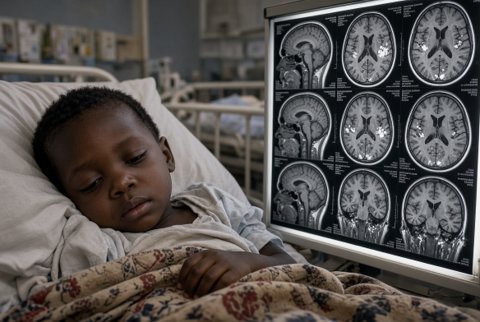
GM1 gangliosidosis is an autosomal recessive lysosomal storage disorder due to biallelic variants in GLB1, with deficiency of beta galactosidase and accumulation of GM1 ganglioside in neurons. It has infantile, late infantile or juvenile, and adult forms.
Pathophysiology. Beta galactosidase removes terminal galactose from GM1 ganglioside, keratan sulphate and oligosaccharides. Deficiency causes neuronal storage and neurodegeneration. Allelic variants in the same gene also cause Morquio syndrome type B (mucopolysaccharidosis IVB), which affects the skeleton without brain disease.
Residual enzyme activity predicts the age of onset.
No disease modifying treatment is established, so care is supportive.
